
One of the problems in creating light emitting diodes (LED) is optimizing their colour properties. Based upon that, organic molecules with heavy metals are used as LED's active material. This particular work is about theoretically investigating the structural and spectral properties of such a molecule – (acetylacetone)bis(2-phenylbenzo[d]thiazole) with iridium (see Fig. 1) – and its constituents.
The goal of this work is to investigate the dependence of the properties of a (pbt)2Ir(acac) complex on the basis function set and on the method of calculation. The work was performed using computational chemistry methods. One of the methods used was the Model Core Potential with large basis function set. Assortment of Density Function Theory functionals was used in calculations as well. All of it was executed on a computation cluster at Physics Faculty of Vilnius University [1]. The results were compared to the earlier theoretical studies and completed experiments.
The results yield that the molecule has three stable conformers. Optimized geometric parameters of the singlet ground state (S0) and lowest triplet state (T1) differ only by a minute amount for the most stable structures. The compared functionals show that it is better to use the B3LYP functional for absorption spectra calculations. Estimated phosphorescence wavelength differs from the experimental results, but may be accepted as the fringe wavelength. Compiled bundles of results of the excited states using S0 and T1 geometries show that the excited states rearrange at different minima (see Fig. 2).
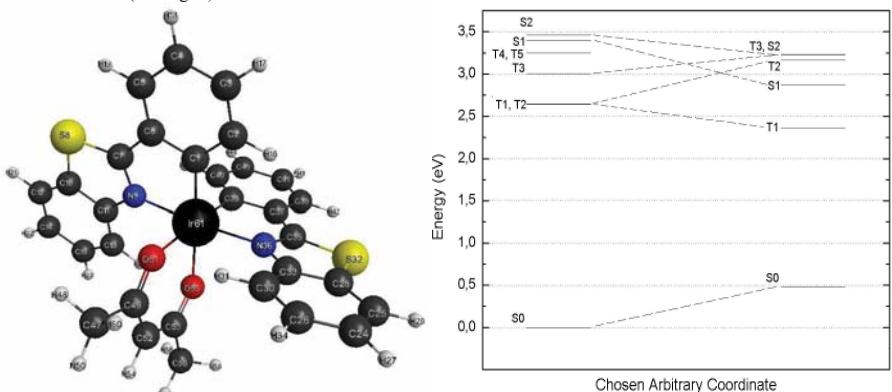
In order to fully explain the results shown in Fig. 2 and to gain an insight on how the molecule's emitting mechanism really works deeper research of the competing transition processes (involving higher electronic states) is needed.