
It is seems that calculation of the exact potential energy curves (PEC) of the diatomic molecules is routine task. But everybody who has dealt with this problem will agree that it is kind of art. Despite a number of papers on the subject [1, 2] we decided to perform a series of calculations.
The potential energy curves (PECs) for the ground and low-lying excited states of KRb are calculated by using high level ab initio MRPT method (XMCQDPT2/CASSCF approximation). The effective core potential (ECP) basis sets for K and Rb atoms are modified ECP10MDF and ECP28MDF respectively. CASSCF (complete active space self-consistent field) and XMCQDPT2 (extended multiconfiguration quasi-degenerate 2nd order perturbation theory) calculations were performed using the Firefly program package. The active space for the CASSCF calculations is 2 electrons in 14 orbitals. All the doubly occupied orbitals in all configurations are active in XMCQDPT2 calculations. To find energy levels values and wave functions the Schrödinger equation for vibrational-rotational motion of diatomic molecule (1) was solved using DVR and Fourier series methods [3,4].
$$-F_{KRb}\frac{d^{2}\Psi}{dx^{2}}+\left\{U(x)+\frac{F_{KRb}J(J+1)}{(x+R_{KRb}^{0})^{2}}\right\}\Psi=E\Psi \tag{1}$$where $x = \frac{R_{KRb} - R_{KRb}^0}{l_0}$ - dimensionless vibrational coordinate, $R_{KRb}$ - distance between atoms, $R_{KRb}^0$ - equilibrium distance between atoms, $l_0$ =1Å, U(x) - PEC, $F_{KRb}$ - kinetic factors depending on the masses of the isotopes of the K and Rb atoms, J – rotational quantum number. Calculated PEC for some exited and ground singlet states are represented on figure 1.
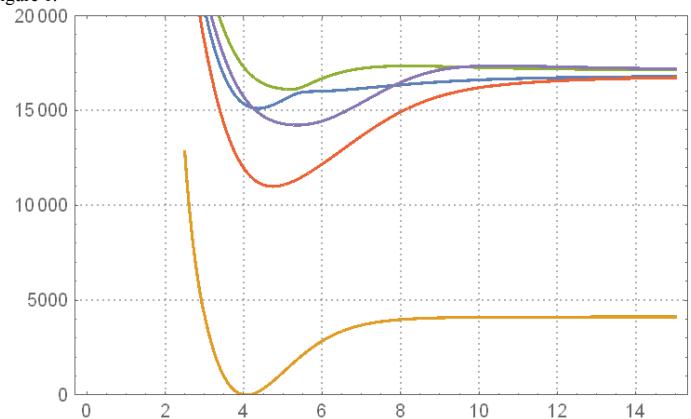
Vibrational-rotational energy levels values for each electronic states and J=0-10 were found. Differences between vibrational level as function of the vibrational quantum numbers for ground state extremely smooth and their values are in perfectly consistent with experimental ones. Spectroscopic constants for different singlet levels were calculated. Franck-Condon factors for transitions from ground vibrational and electronic state to exited electronic and vibrational level were calculated too.