
Ionic liquids (ILs) have been known as “green” and biocompatible solvents for quite some time and their possible biological uses include making molecules permeate more easily through membranes, changing catalytic activity of enzymes, as well as stabilizing or destabilizing various proteins [1]. The nature of the interactions between ILs and proteins varies and is difficult to evaluate. Therefore, smaller peptides with amino acid sequences common in proteins are often used as model systems to discover how different ILs interact with these peptides and change their structure in aqueous solution. Nuclear magnetic resonance (NMR) spectroscopy is the method best suited for identifying both intermolecular interactions and intramolecular structural changes, since the chemical shifts of atoms, particularly hydrogen, are highly sensitive to changes in the local environment of the peptide. 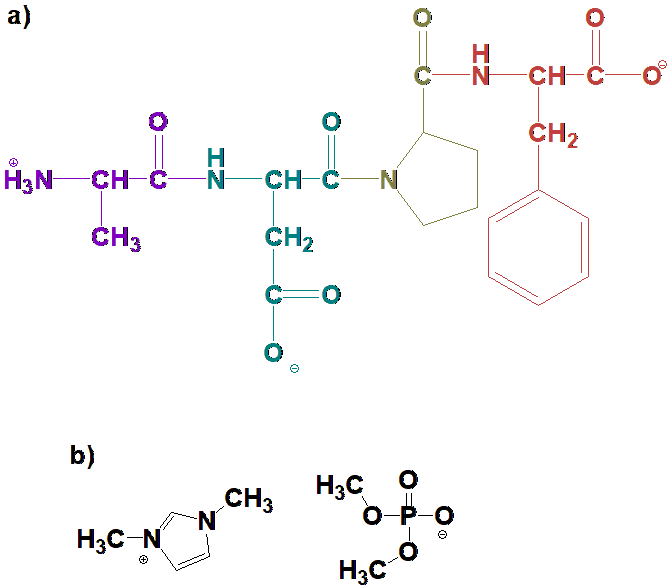
In this work, trans– and cis– Ala-Asp-Pro-Phe tetrapeptides were investigated using molecular dynamics (MD) simulations and quantum mechanics/molecular mechanics (QM/MM) calculations in both aqueous environment and in IL/water mixture. Such systems were analysed experimentally using NMR methods before [2], showing a difference of the peptide’s hydrogen atoms depending on its’ surroundings. The chosen ionic liquid [MMIM][DMP] constituted of 1,3-dimethylimidazolium and a common IL anion dimethyl phosphate. Computed chemical shift differences of atoms enabled us to evaluate various interactions present in peptide/IL/water systems. Meanwhile, trajectories from the MD simulations revealed structural changes in the peptide itself, as well as differences in its immediate surroundings.