
Crystallography allows to determine the exact positions of atoms that constitute crystal structures [1]. Crystal structure determination methods, however, are unable to capture chemical bonding. In practice, two atoms are considered to be connected by a chemical bond if the calculated distance between them is smaller or equal to the sum of their covalent radii. This distance is called a covalent bond length.
Covalent radii tables which are often used to determine chemical bonds between atoms are created by statistically analysing lengths of observed chemical bonds. The most popular covalent radii tables [2] are derived using data from the Cambridge Structural Database [3] which is distributed under a proprietary license and therefore restricts the usage and spread of such derivative datasets. Due to this, there was a clear need to develop a completely independent workflow that allows covalent radii tables to be automatically calculated using open crystallographic datasets such as the Crystallography Open Database [4].
Method used in this research is based on the determination of the van der Waals gap – an interval within the interatomic distance distribution that separates peaks formed by covalent interactions from those formed mainly by van der Waals interactions. This interval contains no interatomic distance observations and thus corresponds to the lowest density region in the distribution [5]. In order to identify the lowest density region, a Gaussian distribution mixture model can be applied to the histogram of interatomic distances.
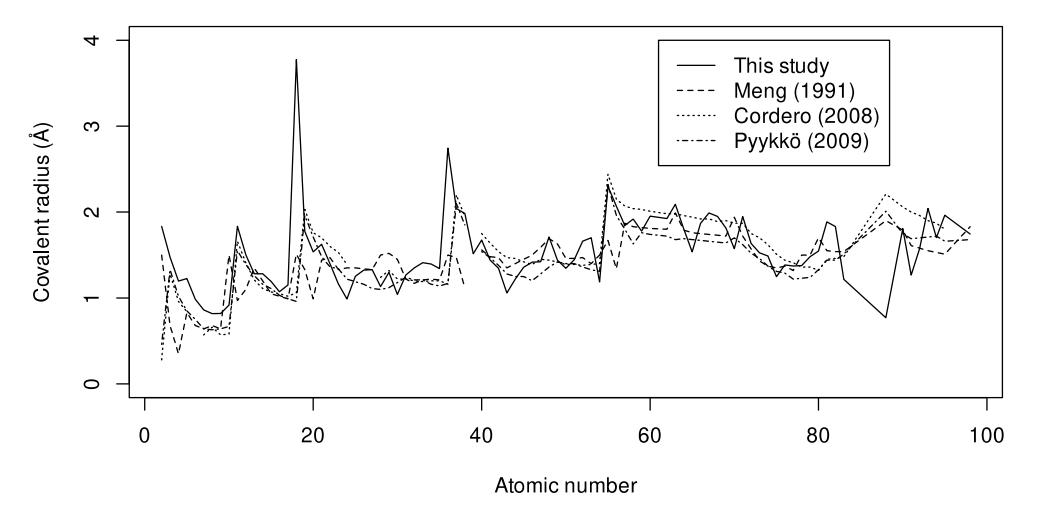
In the current study, a tool for automated determination of covalent radii from interatomic distances was created. Comparison of derived results with selected published covalent radii tables (Figure 1) displays reasonable deviations (up to 0.5 Å) for most atom types. Further research is focused on methods that would help to reduce these deviations that are primarily influenced by the quality of the input data, scarcity of observations in rare atom contact classes and interatomic contacts that involve chemical elements which rarely participate in covalent bonds.
Additional analysis of the input crystallographic data revealed that unmodelled solvent molecules, unmarked disordered structures as well as human errors in crystal structures often lead to abnormal interatomic distance observations. To address this, a set of quality criteria such as compatibility between the declared and calculated properties, use of standard experimental environment conditions, reasonable crystal structure density, etc. were determined to more easily identify crystallographic structures that are suitable for covalent radii derivation calculations. Application and further development of these criteria is part of an ongoing investigation.